- LOGIN
- MemberShip
- 2026-10-11 05:23:06
- Policy
- Preferential drug pricing not enough for emerging innovative pharmas
- by Jung, Heung-Jun Oct 08, 2026 09:00am
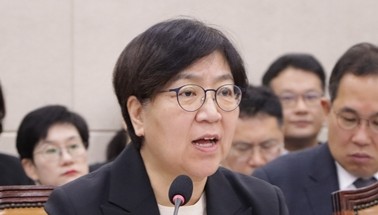
- During a National Assembly audit, calls have emerged for stronger support for “emerging (semi)” innovative pharmaceutical companies, a new intermediate category introduced below innovative pharmaceutical companies.The argument is that preferential drug pricing and government R&D support are not enough to help the semi-innovative companies advance to fully innovative pharmaceutical company status.In response, the Ministry of Health and Welfare (MOHW) said it would gather industry feedback through a public-private consultative body before implementing the new system and review whether additional support measures are needed.During the National Assembly audit of the MOHW on the 7th, People Power Party lawmaker Tae-ho Kim agreed to the government’s policy of dividing innovative pharmaceutical companies into “emerging (semi-innovative)” and “leading (innovative)” categories and providing support according to their stage of growth, but criticized the lack of support offered to the emerging category.Kim said, “I understand the government’s intention to build a growth ladder for pharmaceutical companies by dividing them into emerging and leading categories. But in practice, it seems as if several of the lower rungs of that growth ladder have been removed, then coined ‘emerging’ in name only.”“I’m not sure whether telling companies to jump higher after removing the footholds is really a policy designed to help them grow, or simply a test of how well they can jump,” he added.Support for companies in the emerging category is largely limited to government R&D programs and preferential drug pricing, while excluding benefits available to existing innovative pharmaceutical companies—including tax incentives, special provisions for the construction of research facilities and exemptions from certain charges.Kim said, “We should be helping these companies conduct more research so that they can grow into innovative companies, but this could instead reduce their capacity to invest in research. Can this really serve as a growth ladder?”Health and Welfare Minister Eun-kyeong Jeong responded that the emerging category is being newly introduced as an intermediate step toward full innovative pharmaceutical company status and that support measures would be refined as the program is implemented.Jeong said, “Since this is a new category, we first need to evaluate participating companies and then consider improvements to the system step by step. The emerging category is intended to serve as an intermediate stage toward innovative pharmaceutical company status, with certain forms of support helping companies grow along the way.”Jeong cited preferential treatment under the revised drug pricing system as one of the main reasons for introducing the new category. She nevertheless acknowledged the gap in support between emerging and fully innovative companies and said the government would consider additional measures as the program develops.Jeong said, “I understand that the point is that we need appropriate support measures tailored to different types of companies, creating a pathway from startups to emerging companies and ultimately innovative pharmaceutical companies. We will design the system more carefully and refine it based on industry feedback gathered through the public-private consultative body.”Kim asked the ministry to submit a comparative analysis before the comprehensive audit, including supplementary measures to support clinical development and research funding for emerging innovative pharmaceutical companies.
- Policy
- MFDS cases find new home...but where will rebate probes go?
- by Jung, Heung-Jun Oct 07, 2026 08:52am
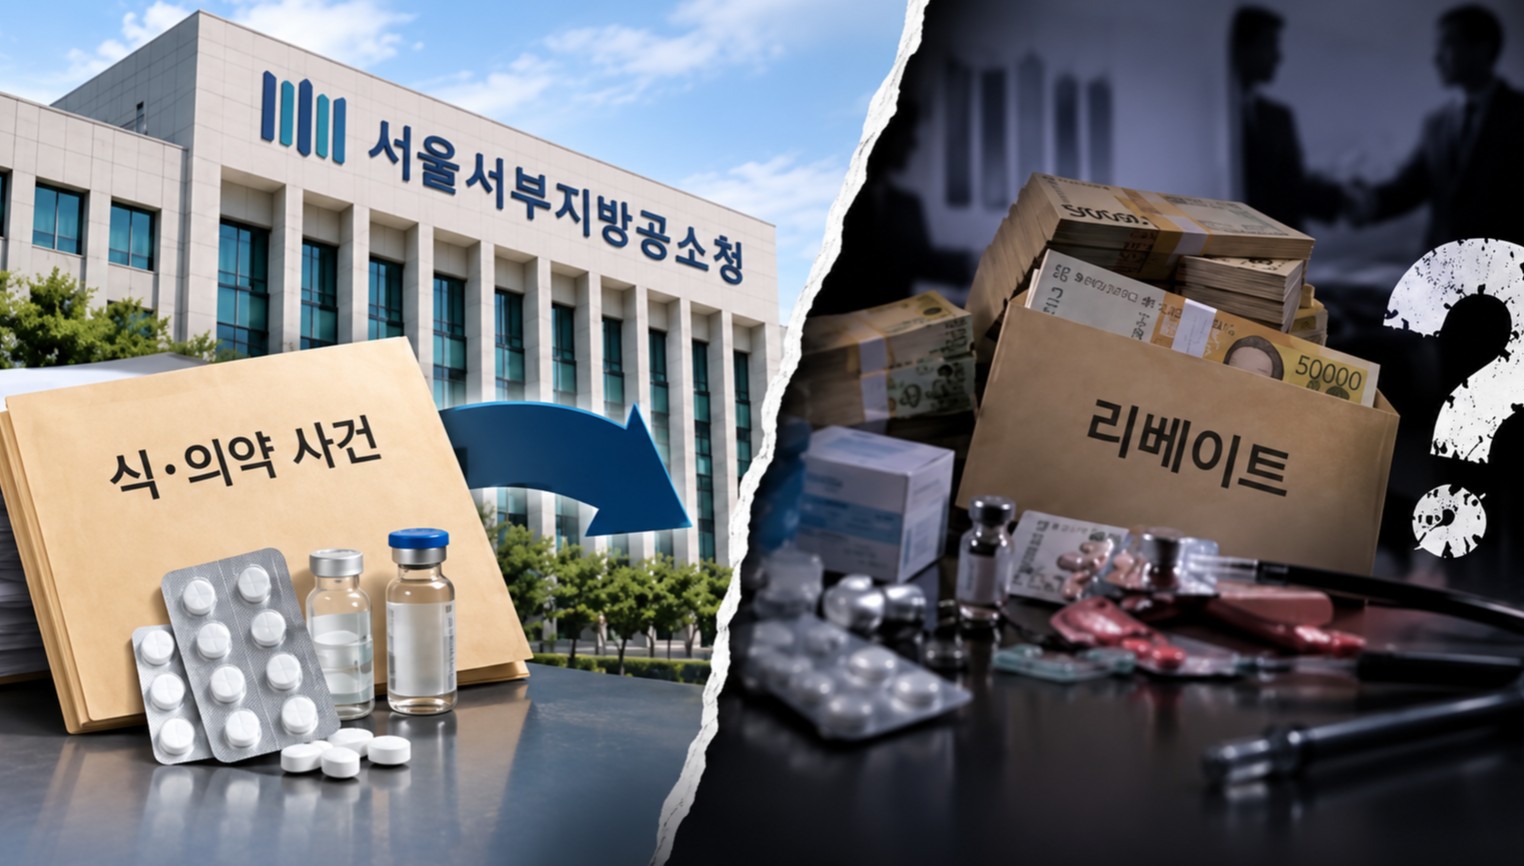
- Food and drug cases previously handled by the Seoul Western District Prosecutors’ Office’s Food and Drug Crime Investigation Division will now largely be handled by the Serious Crimes Prosecution Division of the Seoul Western District Public Prosecution Office. AI-generated image.The investigative framework for Korea’s healthcare sector is taking different paths across different areas following the abolition of the Prosecutors’ Office and the launch of the Public Prosecution Service on October 2.Some ongoing cases have been dispersed to police following the dissolution of joint investigation teams, while in other areas, it remains unclear which body will take over from the specialized investigative units that previously handled them.Food and drug cases investigated by the Ministry of Food and Drug Safety’s Criminal Investigation Office will now be referred to the Seoul Western District Public Prosecution Service instead of the former Seoul Western District Prosecutors' Office..For pharmaceutical and medical device rebate cases, however, the successor investigative authority remains unclear following the dissolution of the Food and Drug Crime Investigation Division of the Seoul Western District Prosecutors’ Office, which had previously led such investigations.A separate joint investigation team focused on illegally operated hospitals and pharmacies operating under borrowed pharmacist licenses has also been disbanded, with its existing cases transferred to different police units.According to relevant agencies on the 6th, the transition to the new prosecution system and dissolution of the Seoul Western District Prosecutors’ Office’s Food and Drug Crime Investigation Division have also reshaped investigative and prosecutorial functions in food and drug cases.The referral system for food and drug cases directly investigated by the MFDS Criminal Investigation Office, however, remains largely unchanged.A Serious Crimes Prosecution Division that has been established within the Seoul Western District Public Prosecution Service will handle food- and drug-related cases. As a result, cases completed by the MFDS unit will continue to be referred to the Seoul Western District Prosecution Service.An MFDS Criminal Investigation Office official said, “The Seoul Western District Public Prosecution Service has established a Serious Crimes Prosecution Division, and its responsibilities include food and drug cases. As before, all cases investigated by our unit will be referred to the Seoul Western District Public Prosecution Service.”The official added, “Since we previously referred all food and drug cases investigated by the unit to the Seoul Western District Prosecutors’ Office, there is little practical change on our part.”The greater uncertainty surrounds investigations into pharmaceutical and medical device rebates. These had largely been conducted through joint investigations led by the Food and Drug Crime Investigation Division of the Seoul Western District Prosecutors’ Office, with police, the Ministry of Health and Welfare, the MFDS and other agencies providing personnel and specialized information under prosecutor-led investigations.The Seoul Western District Public Prosecution Service is still determining how individual cases previously handled by the Food and Drug Crime Investigation Division will be reassigned.An official from the Seoul Western District Public Prosecution Service said. “The work previously handled by the Food and Drug Crime Investigation Division is expected to fall under the Serious Crimes Prosecution Division, but the division of responsibilities has not yet been fully settled.”Regarding where cases from the former investigative areas would be transferred, the official said, “Cases are being handled individually, so it is difficult to provide details on the specific transfers.”The Serious Crimes Prosecution Division is responsible for indictments and maintaining prosecutions in food and drug cases but does not conduct investigations itself.The police can investigate rebate cases, while the Serious Crimes Investigation Agency may also become involved depending on the nature of a case. However, it remains unclear whether the dedicated investigative framework previously led by the Food and Drug Crime Investigation Division will be maintained.The joint investigation team on illegally operated hospitals and license-lending pharmacies, established at the Seoul Western District Prosecutors’ Office in May, was also disbanded ahead of the launch of the new prosecution system.The task force was dissolved as of the 1st, and personnel seconded from agencies including the Ministry of Health and Welfare, the National Health Insurance Service and HIRA have returned to their respective organizations.An official from the task force said, “The team was disbanded as of Oct. 1, and all seconded personnel returned to their original organizations,”.Cases being investigated by the team were also transferred to police. “We handed existing cases over to the police, including some that went to the Seoul Metropolitan Police Agency. Previously, cases were consolidated in one place for investigation, but they have now been split up.”Investigations into illegally operated hospitals and license-lending pharmacies have therefore shifted from a centralized model that brought personnel together in a single task force to a decentralized, police-led system.For areas such as rebates, illegally operated hospitals and license-lending pharmacies, where investigations had been led jointly by prosecutors and other agencies, maintaining specialized investigative capabilities and continuity hereon will be a key challenge under the new system.Meanwhile, the Prosecutors’ Office, which had held both investigative and prosecutorial authority, closed after 78 years, with the Serious Crimes Investigation Agency and Public Prosecution Service launched on the 2nd of this month. The change marks a major restructuring of Korea’s criminal justice system, with investigative and prosecutorial functions formerly exercised by prosecutors now assigned to separate agencies.Under the revised Criminal Procedure Act, Public Prosecution Service Act, and Serious Crimes Investigation Agency Act that took effect on Oct. 2, police are responsible for investigating general criminal cases, while the Serious Crimes Investigation Agency handles serious crimes. The Public Prosecution Service will determine whether to indict cases referred by investigative agencies and conduct prosecutions in court. As a result, criminal complaints and accusations can no longer be filed with the Public Prosecution Service. General cases must be filed with police, while crimes under the jurisdiction of the Serious Crimes Investigation Agency must be filed with that agency.The Serious Crimes Investigation Agency investigates major crimes including corruption, economic crimes, defense procurement offenses, narcotics, national security offenses, cybercrime and abuse of law. It also conducts supplementary investigations into seven categories of crimes involving socially vulnerable victims, including child abuse, domestic violence and sexual offenses. Crimes committed by senior public officials in connection with their duties remain under the jurisdiction of the Corruption Investigation Office for High-ranking Officials.With the abolition of the Prosecutors’ Office, cases previously investigated by prosecutors are also being transferred to the Serious Crimes Investigation Agency or police according to jurisdiction. Prosecutors at the new Public Prosecution Service may, however, continue investigating certain cases for up to 90 days when the statute of limitations is approaching or when consistent handling is required for matters such as warrant applications.
- Policy
- JW Pharma begins developing triple therapy combining Livalozet + SGLT-2 inhibitors
- by Lee, Tak-Sun Oct 07, 2026 08:52am
- JW Pharmaceutical has officially begun developing a next-generation fixed-dose combination (FDC) triple therapy, combining an SGLT-2 inhibitor-class antidiabetic agent into its dyslipidemia treatment, 'Livalo.'The company's strategy aims to solidify its leadership in the chronic multimorbidity market by merging the potent lipid-lowering efficacy of the existing 'Livalozet (pitavastatin + ezetimibe)' with glycemic control.According to the Ministry of Food and Drug Safety (MFDS), JW Pharmaceutical secured Investigational New Drug (IND) approval on the 2nd for a Phase 1 clinical trial of its investigational FDC triple therapy candidate, 'JW0113.'This clinical study will enroll 64 healthy adult volunteers and is scheduled to run from December 2026 to December 2027. The trial is designed as an open-label, randomized, 2-sequence, 4-period, fasting, single-dose, oral, replicated crossover study. The objective is to demonstrate bioequivalence by comparing the pharmacokinetic (PK) profiles and safety of the FDC candidate 'JW0113' administered alone versus co-administration with the corresponding reference listed drugs.Notably, as the primary pharmacokinetic endpoints specify the area under the plasma concentration-time curve from time zero to the last measurable concentration (AUCt) and peak plasma concentration (Cmax) for pitavastatin, total ezetimibe, and empagliflozin, it was confirmed that this pipeline candidate is an FDC triple therapy combining the active pharmaceutical ingredients of Livalozet with empagliflozin, a flagship SGLT-2 inhibitor.The chronic disease FDC market pioneered by 'Zemiglo' and 'Rosumet'… now advancing to 'triple combination therapy'With more than 70% to 80% of domestic diabetic patients concurrently presenting with dyslipidemia, the two conditions represent a quintessential multimorbidity. Due to the clinical nature of these diseases, patients often face a high pill burden, which tends to compromise medication adherence. Consequently, the pharmaceutical industry moved early to introduce combination therapies to manage both conditions simultaneously.In the domestic market, products such as LG Chem's 'Zemiglo (gemigliptin + rosuvastatin, a DPP-4 inhibitor + statin)' and Yuhan's 'Rosumet (rosuvastatin + metformin, a statin + first-line antidiabetic agent)' opened the era of 'diabetes + dyslipidemia' FDCs and are recognized as benchmark cases that enhanced patient dosing convenience.However, as recent cardiovascular disease prevention guidelines have adopted more stringent LDL cholesterol targets, the mainstream paradigm in lipid management has shifted from 'statin monotherapy' to 'statin + ezetimibe combination therapy.' Concurrently, combination regimens involving 'SGLT-2 inhibitors,' which have demonstrated cardiovascular and renal protective benefits, have become the standard of care in diabetes management. Accordingly, as market demand moves beyond existing dual therapies (one statin + one antidiabetic agent), there is rapid emerging demand for a generational transition to 'triple FDCs' that fuse the most potent lipid-lowering combination, 'statin + ezetimibe', with next-generation antidiabetic agents.Evolution of the 'Livalo Family'… Expanding from hypertension and mixed dyslipidemia into 'diabetes'JW Pharmaceutical has continuously diversified its product portfolio, anchored by its pitavastatin monotherapy, 'Livalo Tab.' The company has established a comprehensive cardio-cerebrovascular and chronic disease therapeutic portfolio, spanning 'Livalo V (pitavastatin + valsartan)', the statin + ezetimibe combination 'Livalozet' introduced in 2021, the fenofibric acid combination 'Livalo Feno' approved in 2025, and the triple-combination antihypertensive 'Livalo Hi (valsartan/amlodipine/pitavastatin).'The development of "JW0113" is regarded as a pivotal strategic bridge linking the Livalo family directly to the "diabetes space."In particular, empagliflozin is a benchmark SGLT-2 inhibitor with established efficacy not only in robust glycemic reduction but also in lowering the risk of heart failure and renal complications. Furthermore, compared to other statins, pitavastatin demonstrates a lower risk of new-onset diabetes mellitus (NODM), driving strong prescribing preferences for diabetic patients or individuals at high risk of diabetes. By combining these two agents with ezetimibe, which inhibits intestinal cholesterol absorption, the regimen will provide an optimal therapeutic option in a single pill for patients who have concurrent dyslipidemia and diabetes.A pharmaceutical industry official said, "While conventional dual-combination therapies like Zemiglo and Rosumet improved dosing convenience, JW0113 is an evolved model that brings three key drugs aligned with the latest clinical treatment guidelines into a single pill," and added, "If JW Pharmaceutical successfully commercializes this triple FDC alongside Livalo in the market, it is expected to secure an unmatched competitive edge across the chronic disease sector."
- Policy
- MOHW opposes bill requiring proof for CSO registration
- by Lee, Jeong-Hwan Oct 07, 2026 08:52am
- The Ministry of Health and Welfare (MOHW) has effectively opposed a proposed amendment to the Pharmaceutical Affairs Act that would explicitly stipulate in law the requirements for CSOs to maintain a place of business and require the submission of supporting documents, arguing that the change would offer little practical administrative benefit.On the 2nd, the ministry was found to have submitted its position on the bill proposed by Democratic Party lawmaker Jin-sook Jeon, which is pending in the National Assembly.Jeon’s bill would require CSOs, when reporting their pharmaceutical promotional sales activities to local governments, to meet requirements prescribed by Ministry of Health and Welfare regulations, including maintaining at least one business premises, and submit documents demonstrating compliance. The measure is intended to prevent the proliferation of so-called “ghost CSOs” that operate without an actual place of business.While agreeing with the bill’s aim of improving transparency in pharmaceutical distribution, the ministry stopped short of supporting the legislative change, citing its limited practical effect. Its position is that the existing framework already provides sufficient means to verify and enforce the requirements, reducing the need for a statutory amendment.The MOHW explained, “The current Enforcement Rule of the Pharmaceutical Affairs Act already stipulates that a CSO must have a business address as a reporting requirement, and local governments do not accept reports if the address cannot be verified.”“Competent authorities can already sufficiently verify the existence of an actual place of business through official documents submitted with the report, such as a business registration certificate or corporate registry. Since the requirements can already be enforced under the current rules, there is little practical benefit to amending the law.”The National Assembly’s Expert Advisory Office also lent support to the ministry’s position, noting that while elevating provisions currently contained in subordinate regulations into the parent statute could improve legal clarity, the practical benefit of the amendment would be limited because compliance can already be verified through checklists and official documents.The advisory office further noted that simply verifying the existence of business premises would do little to prevent illegal or evasive practices by disguised businesses, and suggested that more substantive measures are needed, such as mechanisms to identify ownership structures or new authority to investigate pharmaceutical sales and distribution practices.The Korean Pharmaceutical Association (KPA), however, strongly supported the amendment, calling for its swift passage to address problems associated with ghost CSOs.KPA said, “The recent increase in CSOs operating without business premises has made pharmaceutical distribution channels and lines of accountability increasingly unclear, worsening blind spots in oversight and contributing to disruptions in distribution practices, including illegal rebates and unfair sales practices. The amendment would clearly establish in law the basic infrastructure and reporting requirements for pharmaceutical sales representatives, thereby improving transparency in distribution channels and strengthening accountability.”
- Policy
- MFDS moves to speed finished drug approvals
- by Lee, Tak-Sun Oct 06, 2026 08:48am
- The Ministry of Food and Drug Safety (MFDS) is changing how it communicates requests for supplementary information on active pharmaceutical ingredients (APIs) during the approval process for finished pharmaceutical products, sending them directly to API manufacturers rather than through finished-drug companies.The change is expected to reduce review delays and improve technical communication by eliminating the multi-step process in which reviews have traditionally been delayed at the finished-drug company stage.The MFDS Drug Approval Policy Division said on the 30th that it will launch a pilot program, starting with applications for new or amended approvals filed from Oct. 1, under which API manufacturers will be directly notified of requests for supplementary information arising from linked reviews of APIs and finished products.‘Too much time lost going through finished-drug companies’...addresses industry communication difficultiesThe linked ‘API-finished product review’ is a system under which the quality and safety of the active ingredient registered under the Drug Master File (DMF) system are reviewed in conjunction with the marketing authorization application for the finished pharmaceutical product.Problems first arose when technical questions or requests for additional information concerning the API emerge during the review. Because the finished-drug company is the formal applicant, the MFDS has traditionally sent API-related requests only to that company, which then has to forward them to the API manufacturer and relay its response back to the agency.The industry has repeatedly raised concerns that this multi-step process can distort highly technical questions involving areas such as synthesis processes and impurity control. Companies have also reported difficulty meeting submission deadlines when API manufacturers are reluctant to share sensitive trade secrets with finished-drug companies.New drugs and data-submission drugs first...pilot limited to domestic API manufacturersThe key change is that the MFDS will notify API manufacturers directly by official letter, allowing the companies that actually generate the data and possess the relevant technical expertise to immediately identify and respond to requests for supplementary information.The pilot will cover new drugs, data-submission drugs, and follow-on drugs (generics), as well as applications for changes to their approvals handled by the Pharmaceutical Approval Management Division. It will apply when a finished-drug applicant submits a data-sharing consent form from a DMF holder (Korean API company, including the original registrant and APIs registered through a letter of authorization).The MFDS will send the relevant API manufacturer an official letter specifying the API name and DMF registration number, along with requests for “supplementary information related to Section 3.2.S, Drug Substance,” of the Common Technical Document (CTD).The pilot will not apply to products for which a data-sharing consent form has been submitted by an overseas API manufacturer, or where the DMF holder and finished-drug company are the same legal entity. Products under the jurisdiction of regional food and drug safety offices are also outside the scope of the current pilot.An MFDS official said, “By sending requests for supplementary API information directly to the API manufacturers responsible for preparing the data, we expect to improve review efficiency and support faster approval. We will closely assess the results of the pilot program before deciding how long to continue it and whether to expand its scope.”
- Policy
- Drug pricing negotiations are underway for Ofev·Lynparza
- by Jung, Heung-Jun Oct 06, 2026 08:48am
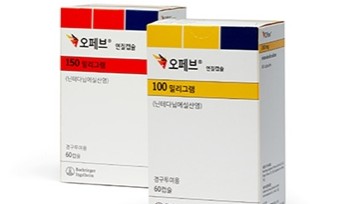
- Product photo of Ofev Soft CapsulesBoehringer Ingelheim's Ofev Soft Capsules (nintedanib esylate) and AstraZeneca's Lynparza Tablets (olaparib) are currently undergoing drug price negotiations, the final stage before reimbursement listing.Furthermore, Ipsen Korea's Cabometyx Tablet (cabozantinib) and Kyowa Kirin Korea's Crysvita Solution for Injection (burosumab), both recognized for the appropriateness of their reimbursement expansion at the Pharmaceutical Reimbursement Evaluation Committee (PREC) meeting last July, are also in the process of negotiation.According to industry sources on the 2nd, drug price negotiations are underway for Ofev Soft Capsules, for which an application to expand the reimbursement scope to include idiopathic pulmonary fibrosis (IPF) was submitted last year.Ofev, which received domestic approval in 2016, was added to the reimbursement listing in May of last year after approximately nine years of persistent efforts. At that time, national health insurance coverage was applied for ▲ systemic sclerosis-associated interstitial lung disease (SSc-ILD) and ▲ progressive pulmonary fibrosis.However, because Ofev failed to obtain reimbursement approval for idiopathic pulmonary fibrosis, the company submitted an application for reimbursement expansion last year.The company has dedicated significant effort to this reimbursement expansion, including submitting real-world data (RWD) on patients who failed first-line treatment with 'Pirespa' (pirfenidone) (including treatment discontinuation due to adverse events).Patient advocacy groups, including the Korean Organization for Rare Diseases, have also urged reimbursement expansion, noting that South Korea is the only country among developed nations, including the A8 countries, where insurance coverage does not apply to Ofev for idiopathic pulmonary fibrosis.Having reached the drug price negotiation table 10 years after its domestic approval, the reimbursement expansion for idiopathic pulmonary fibrosis is now imminent.Three additional products are currently under negotiation for reimbursement expansion: Lynparza, Cabometyx, and Crysvita.AstraZeneca's Lynparza was recognized by the PREC in July for reimbursement expansion as a maintenance therapy in combination with bevacizumab for adult patients with ovarian cancer, fallopian tube cancer, or primary peritoneal cancer.Additionally, Ipsen Korea's Cabometyx was determined appropriate for reimbursement expansion as a monotherapy for metastatic or recurrent clear cell renal cell carcinoma in patients who have failed first-line treatment, and Kyowa Kirin Korea's Crysvita received a decision on appropriateness for adult patients with FGF23-related hypophosphatemic rickets and osteomalacia.Because the negotiation orders for these drugs were issued in August and September, these products are scheduled to be listed as early as November or December.
- Policy
- Rybrevant reimbursed under triple RSA agreement in Korea
- by Jung, Heung-Jun Oct 02, 2026 10:23am
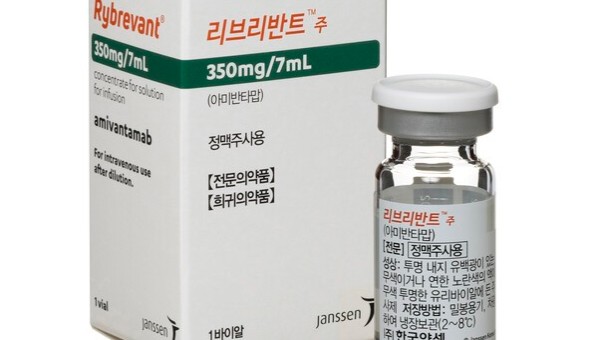
- Janssen Korea’s non-small cell lung cancer treatment Rybrevant (amivantamab), which had sought reimbursement for more than 4 years, is understood to have secured coverage under a three-part risk-sharing agreement (RSA).In addition to refund and expenditure-cap arrangements, the agreement includes post-listing management based on the collection of real-world data (RWD). The approach is intended to reassess clinical uncertainty after listing using data generated in real-world practice.According to industry sources on the 1st, Rybrevant, which gained reimbursement this month, is subject to a combination of a refund-based RSA, an expenditure cap, and post-listing real-world data collection.Rybrevant was approved by the Ministry of Food and Drug Safety in 2022 and gained reimbursement this month after several attempts over roughly 4 years.After applying for reimbursement in the year of its approval, the drug failed on multiple occasions to establish reimbursement criteria at the Cancer Disease Review Committee level. Its path to listing remained difficult this year, with the Drug Reimbursement Evaluation Committee deciding in May that the drug required further reviewThe indication reimbursed this month is Rybrevant as ‘monotherapy for adult patients with locally advanced or metastatic non-small cell lung cancer with EGFR exon 20 insertion mutations whose disease has progressed on or after platinum-based chemotherapy.’The combined RSA appears to have played an important role in Rybrevant clearing the reimbursement hurdle. With the drug entering into a refund-based agreement, the number of active ingredients covered by refund-based RSAs has risen to 74 as of this month.The agreement also incorporates an expenditure cap and real-world evidence collection. Rather than simply refunding a portion of drug expenditures, the arrangement provides for a drug-level outcomes assessment using real-world data to address limitations in the available clinical evidence.Rybrevant is an oncology drug for a small patient population, with fewer than 200 eligible patients annually identified during price negotiations.The annual treatment cost is approximately KRW 80 million per patient. With the 5% coinsurance rate applied to cancer patients, the patient’s annual out-of-pocket cost is approximately KRW 4 million.Rybrevant’s listed price is KRW 1,322,570. Projected claims expenditure is approximately KRW 12.5 billion, but actual National Health Insurance spending is expected to be lower because both the refund and expenditure-cap arrangements apply.
- Policy
- Amendment to the Pharmaceutical Affairs Act passes NA
- by Lee, Jeong-Hwan Oct 02, 2026 10:22am
- An amendment to the Pharmaceutical Affairs Act strengthening the investigative and rejection authority of local governments over pharmacy registrations passed the National Assembly plenary session on the 1st, tightening regulations on controversial warehouse-style pharmacies and illegally operated pharmacies involving license lending.The amendment also introduces an intermediate sanction allowing GMP certification to be ‘suspended for up to 6 months,’ as part of efforts to improve the current one-strike system under which GMP violations can result in revocation regardless of their severity.The National Assembly held a plenary session and passed the amendment that was consolidated and coordinated by the Health and Welfare Committee. The revised law will take effect 6 months after promulgation.Under the supplementary provisions, however, the new rules governing registration of illegal pharmacies will apply only to pharmacy registration applications submitted to local governments after the law takes effect. Regarding the provision on plans to provide rare disease drugs free of charge, products already being supplied following review by the MFDS commissioner will be deemed approved without additional procedures after the revised law takes effect.Local governments gain authority to ‘request documents, reject pharmacy registrations’… bars entry of warehouse-style, illegally operated pharmaciesThe amendment strengthens pre-registration scrutiny by giving municipal and district authorities statutory authority to request documents needed to determine whether a proposed pharmacy operation would violate the law.Under the current Pharmaceutical Affairs Act, local governments have had difficulty refusing registration when an application formally satisfies registration requirements, even when they need to determine in advance whether the pharmacy involves license lending, collusion with a medical institution, or violations of location restrictions.As a result, authorities have lacked the power to demand detailed supporting documents when applications are filed for warehouse-style pharmacies modeled after large retail stores or logistics warehouses, or for pharmacies effectively owned by non-pharmacists through license-lending arrangements, leaving them unable to reject such applications on those grounds.Under the amended law, mayors, county heads and district heads may, before accepting a pharmacy registration, require the applicant pharmacist to submit documents prescribed by Ministry of Health and Welfare regulations to verify whether the pharmacy ▲violates restrictions on establishment (including operation within the premises or facilities of a medical institution); ▲involves the lending of a pharmacist license; or ▲violates the principle that one pharmacist may establish and operate only one pharmacy.The law also explicitly allows local authorities to reject a registration application if the applicant fails, without legitimate reason, to supplement or submit the requested documents. The same document-request and rejection provisions will apply when a change in ownership of an existing pharmacy requires notification of succession to the status of the pharmacy operator.Up to ‘6-month GMP suspension ’ introduced...sanctions made more proportionateThe GMP certification sanctions system, which garnered major attention from the pharmaceutical industry, has also been revised with the introduction of intermediate penalties.Under the current law, repeated false or incorrect preparation of manufacturing and quality-control records after obtaining GMP certification can result in mandatory revocation of the certification under the ‘one-strike out’ system. The system has faced criticism for imposing potentially severe consequences on businesses and threatening the supply of essential medicines regardless of the seriousness of the violation.Critics have pointed out that manufacturing and quality-control records are extensive and varied, yet violations have been subject to the same uniform revocation penalty without regard to their severity.The amended law introduces intermediate sanctions allowing the MFDS commissioner, depending on the nature of the violation, to suspend the validity of GMP certification for ‘up to 6 months’ or issue a corrective order.For serious violations, however, the amended law retains the strict principle of mandatory revocation, while clearly defining which types of violations warrant mandatory revocation and which are subject to other sanctions.Mandatory revocation applies to cases where GMP certification was obtained through false or other improper means (Item 1), and where a manufacturer repeatedly failed to prepare or retain batch-specific manufacturing and quality-control records and sold the drugs concerned (Item 2).Cases subject to suspension, revocation or corrective orders include manufacturing a drug differently from its approved or notified specifications while falsifying records to make it appear that the drug was manufactured in compliance with those specifications (Item 3), as well as other repeated falsification or erroneous preparation of manufacturing and quality-control records (Item 4).The amendment also includes a safeguard for drug supply. Even when a product’s GMP certification has been suspended, the MFDS commissioner may temporarily permit its manufacture and sale for a specified period if the drug is essential for patient treatment and no alternative is available.Free supply of rare disease drugs legalized...Korea Orphan & Essential Drug Center to oversee programsExpanded access programs (EAPs) providing rare disease drugs free of charge will also be formally brought within the legal framework.Until now, multinational pharmaceutical companies seeking to provide rare disease drugs to patients free of charge on compassionate grounds before reimbursement listing have faced regulatory limitations because such supply could potentially fall under the Pharmaceutical Affairs Act’s prohibition on providing economic benefits (rebates) to medical institutions.The amendment explicitly recognizes the free supply of rare disease drugs by pharmaceutical companies to medical institutions as a permitted economic benefit (exception to the rebate prohibition provision).A pharmaceutical company seeking to provide a rare disease drug free of charge as part of a patient support program must first obtain approval of the plan from the MFDS commissioner, while the Korea Orphan & Essential Drug Center will oversee distribution. After obtaining the patient’s consent, a medical institution will submit an application to the center. The center will then verify whether the request complies with the approved free-supply plan before authorizing the pharmaceutical company to provide the drug. Transitional provisions also stipulate that free-supply programs already underway upon MFDS review before promulgation of the amendment will be deemed to have received approval under the revised law.
- Policy
- Impurity standards for antibiotics relaxed…confusing product recalls
- by Lee, Tak-Sun Oct 01, 2026 09:36am
- AI 생성 이미지The pharmaceutical industry's concerns have eased as nitrosamine impurities detected in macrolide antibiotic active pharmaceutical ingredients have been classified as "non-mutagenic." However, because health authorities finalized their safety reviews after the testing data submission deadlines, pharmaceutical companies have repeatedly faced administrative confusion, voluntarily recalling products only to reverse the decisions, prompting calls to strengthen the regulatory framework.The current domestic and international impurity management system underwent a major shift after the 2018 incidents involving the hypertension drug 'valsartan' and the 2019 gastrointestinal drug 'ranitidine,' in which nitrosamine impurities, including the probable human carcinogen NDMA (N-nitrosodimethylamine), were detected.Moving away from universal regulations and total sales bans, an ongoing management system in which pharmaceutical manufacturers independently evaluate and test risks and voluntarily decide whether to initiate recalls when limits are exceeded has become the global standard.Under this self-regulatory system, the Ministry of Food and Drug Safety (MFDS) recently classified 'N-nitroso-N-desmethyl roxithromycin,' an impurity in roxithromycin, as non-mutagenic, easing concerns for pharmaceutical companies marketing this active ingredient. This follows the July recognition that clarithromycin's impurity, 'N-nitroso-N-desmethyl clarithromycin,' is non-mutagenic."Non-Mutagenic" with no genetic alteration risk... Relaxed to the level of general organic impurities"Non-mutagenic" means that the impurity poses no risk of damaging human cellular genetic material (DNA) or inducing mutations.Under current global standards, for 'mutagenic (genotoxic)' impurities carrying carcinogenic risks, pharmaceutical companies must preemptively execute voluntary recalls even if traces exceed a threshold of merely tens of nanograms (ng) per day. Conversely, if additional mutagenicity testing proves an impurity carries no genetic risk and confirms it as "non-mutagenic," it falls under the general organic impurity management standards of the International Council for Harmonization of Technical Requirements for Pharmaceuticals for Human Use (ICH Q3A/B). As acceptable limits are relaxed by hundreds of times or more, products are effectively freed from quality non-compliance or recall risks.Safety cases continue across macrolide antibiotics as a whole. Previously, 'N-nitroso-N-desmethyl-azithromycin' and 'N-nitroso-azaerythromycin' in azithromycin were recognized as non-mutagenic, and genotoxicity concerns have now been cleared sequentially for impurities in clarithromycin and roxithromycin as well.The problem arises within a framework where pharmaceutical manufacturers must independently judge whether to conduct recalls based on their own testing results. Yet, the regulatory authorities' final review schedules for non-mutagenicity exceed data submission deadlines. Pharmaceutical companies that preemptively decided on voluntary recalls under strict, existing interim limits repeatedly cancel recalls once the non-mutagenicity of the impurities is belatedly confirmed.In fact, JW Shinyak's clarithromycin product, "Lithromycin Dry Syrup 125mg/5mL," initially reported a voluntary recall for exceeding the existing limits, but canceled the recall and resumed normal distribution immediately after the non-mutagenicity determination.Roxithromycin formulations are following the same footsteps. SamaPharm recently announced a voluntary recall for certain lot numbers of 'Roxithromycin Tab. Sama' currently in distribution; however, after confirming the impurity as non-mutagenic, the company is reportedly actively considering retracting the recall, similar to JW Shinyak."Reflecting the purpose of the self-regulatory system"... Administrative flexibility and policy improvements neededIndustry voices together that policy flexibility is required during the review process to preserve the system's core purpose of voluntary quality control and preemptive action by pharmaceutical companies. If premature voluntary recalls take place while impurity safety reviews are still underway, public announcements alone undermine product credibility, creating substantial societal costs from returns and restocking across clinics, hospitals, pharmacies, and wholesale distributors.An official from the pharmaceutical industry stated, "While an advanced system where pharmaceutical companies manage risks on their own has been established since the valsartan crisis, currently, verifying the mutagenicity of an impurity takes considerable time," adding, "For substances currently undergoing safety reviews, urgent administrative policy improvements are needed, such as deferring data submission or voluntary action deadlines until the final results are released."
- Policy
- Retevmo, Vyloy and Padcev+Keytruda reimbursed from Oct. 1
- by Lee, Jeong-Hwan Oct 01, 2026 09:36am
- National Health Insurance reimbursement will take effect on Oct. 1 for Retevmo Cap, a treatment for non-small cell lung cancer with a rare genetic alteration and medullary thyroid cancer; Vyloy, a treatment for advanced or metastatic gastric cancer; and the Padcev-Keytruda combination as first-line treatment for urothelial cancer.The expanded reimbursement coverage is expected to help extend the lives of patients with rare and underserved cancers who have faced high out-of-pocket drug costs, while easing the financial burden on their families.On the 30th, the Ministry of Health and Welfare said that the Health Insurance Policy Deliberation Committee had approved expanding NHI coverage for the cancer treatments.Retevmo Cap is a targeted therapy for non-small cell lung cancer and medullary thyroid cancer with a rare genetic alteration (RET fusion). The number of eligible patients in Korea is estimated at fewer than 300. Compared with conventional cytotoxic chemotherapy, the drug reduces adverse effects while improving treatment efficacy. As an oral therapy, it also substantially reduces the burden of hospital visits and waiting times for patients.Vyloy is a new treatment for patients with unresectable advanced or metastatic gastric cancer, a difficult-to-treat disease with a 5-year relative survival rate of around 10%.Compared with existing treatment, Vyloy was shown to reduce the risk of developing resistance by 41% and nearly double overall survival.NHI coverage will also apply to the Padcev-Keytruda combination as first-line treatment for urothelial cancer. The combination will be covered from the first line of treatment for patients with urothelial cancer, which accounts for approximately 2.5% of all cancers. Combining Padcev, which targets cancer cells, with Keytruda, which activates immune cells, nearly doubled survival compared with conventional treatment.Notably, reimbursement of the Padcev(Astellas Korea) and Keytruda (MSD) combination, respectively, marks the first time Korea’s NHI has covered a combination regimen involving high-cost new drugs developed by different pharmaceutical companies. The case also carries policy significance as coverage was achieved by coordinating separate negotiations and interests between the two drugmakers to close a gap in treatment access. The government plans to use the case as a basis for further improvements to the NHI reimbursement framework for combination regimens involving new drugs.Hyung Hoon Lee, Second Vice Minister of Health and Welfare, said, “This will substantially ease the financial burden on patients and families who have had to delay or forgo treatment because of high drug costs. We will continue endeavors to expand National Health Insurance coverage to treatments for rare and severe diseases.”